Monitoring system evolution through machine-learning molecular dynamics
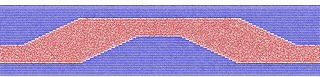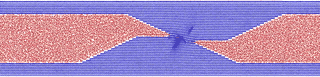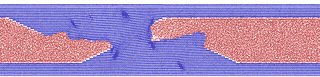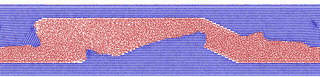
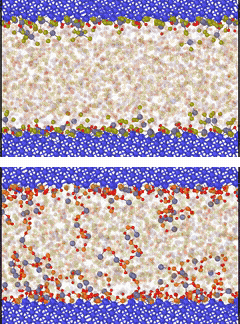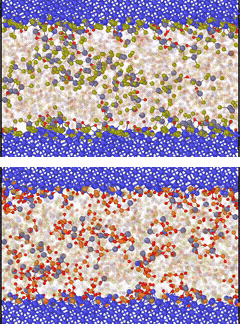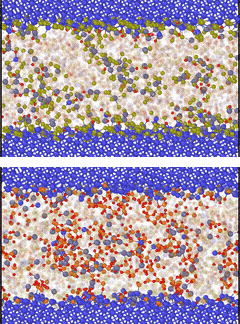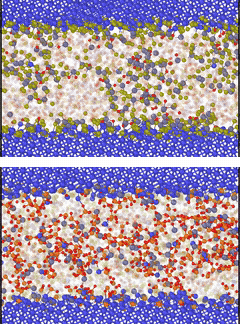
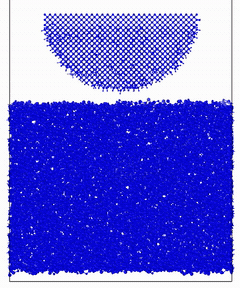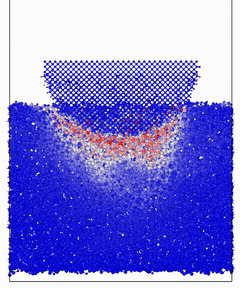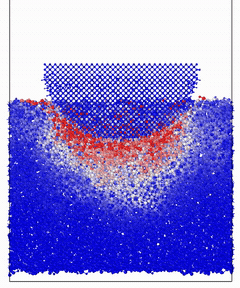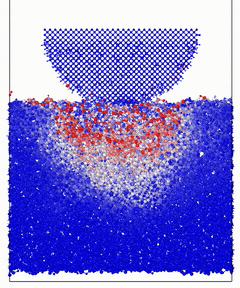
Simulating surface and interface processes for technological applications requires length and time scales beyond the reach of ab initio molecular dynamics. With the advent of machine learning interatomic potentials (MLIPs), it has become possible to perform simulations with near-DFT accuracy at a computational cost comparable to that of classical force fields. We pioneered their use in computational tribology [98, 100] and developed SCS, a software package that automates the construction of training databases through active learning and ad hoc system collaging for tribological applications [109].
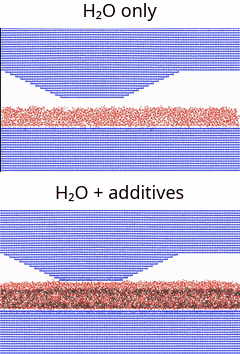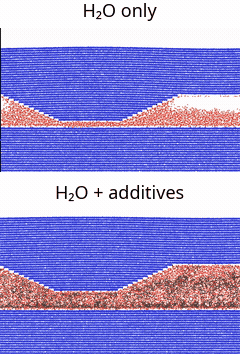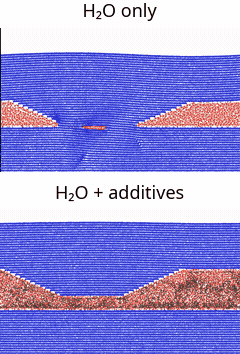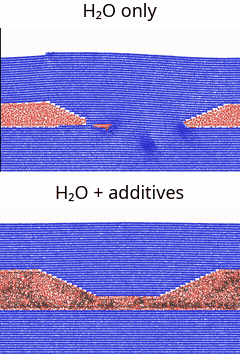
Thanks to these advances, we are able to observe tribofilm formation at nanoasperity contacts in real time. For example, we have unravelled key atomistic mechanisms underlying the action of ZDDP [130], a widely used antiwear additive studied for decades, and related the frictional behavior of different additives to their molecular structure [107, 133]. Ongoing studies explore how carbon-based molecules form low-friction tribofilms and improve water-based lubrication, offering greener alternatives to petroleum-based oils.
MLIPs also allow us to investigate atom by atom wear, simulating nanometer-scale diamond asperities plowing across hard surfaces [117].
High-throughput screening of surface and interface properties
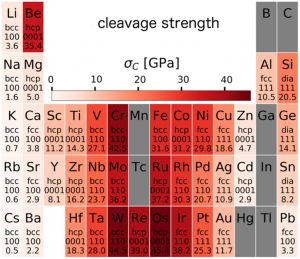
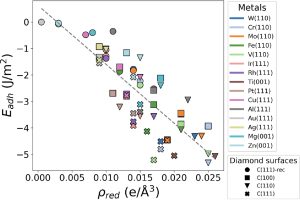
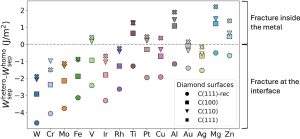
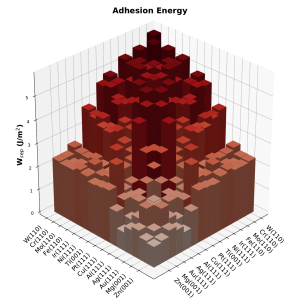
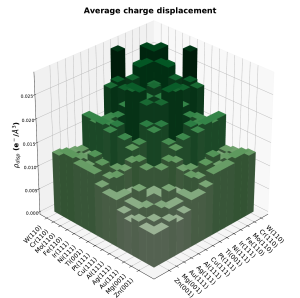
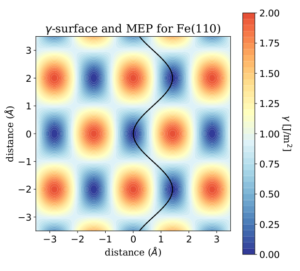
The use of first-principles calculations based on density functional theory (DFT) to screen the properties of many materials in parallel and in an automatized way represents an exciting opportunity for discovering new materials and/or general trends.
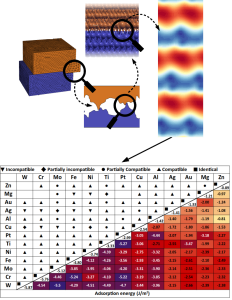
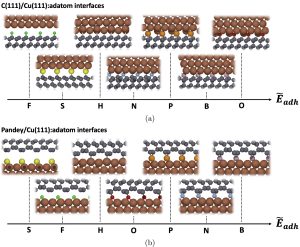
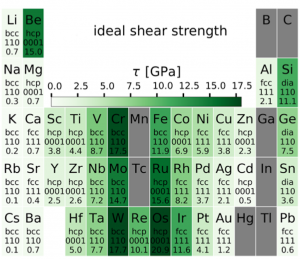
We developed workflows, XSORB and Tribchem, for the automatized screening of solid surfaces and interfaces. The calculated properties include surface adsorption, interfacial adhesion, charge displacements, and ideal shear strength.[47, 53] The latter is obtained from the first-principles calculated potential energy surface (PES) (or Generalized Stacking Fault Energy Surface) according to the procedure developed by our group [23].
Through this approach we calculated the adhesion energies of several metallic interfaces [82], and analyzed how chemical surface modifications affect them [96]. Metal-semiconductor [101, 126], 2D [34, 74], 2D-metal [77] and other classes of solid interfaces relevant to technological applications are under investigation.
The databases we are continuously populating are available here.